Journal of Tropical Diseases and Parasitology ›› 2026, Vol. 24 ›› Issue (4): 236-240.doi: 10.20199/j.issn.1672-2302.2026.04.009
• EXPERIMENTAL STUDY • Previous Articles Next Articles
WANG Yuanyuan1( ), ZUO Yancai1, LIU Xiaosong1, QIU Shenwei1, HUANG Li2, CUI Hongjuan3, PAN Kai3
Received:2026-04-29
Online:2026-08-20
Published:2026-09-29
CLC Number:
WANG Yuanyuan, ZUO Yancai, LIU Xiaosong, QIU Shenwei, HUANG Li, CUI Hongjuan, PAN Kai. Whole-genome sequencing and genetic characterization of norovirus from diarrhea patients in Tongling City[J]. Journal of Tropical Diseases and Parasitology, 2026, 24(4): 236-240.
Add to citation manager EndNote|Ris|BibTeX
URL: http://www.rdbzz.com/EN/10.20199/j.issn.1672-2302.2026.04.009
Table 1
Distribution of genomic regions of the three NoV whole-genome sequences(nt)
| 序列区域 | NR18 | NR61 | NR246 |
|---|---|---|---|
| 全长 | 7 641 | 7 784 | 7 567 |
| ORF1 | 1~5 319 | 1~5 371 | 1~5 095 |
| ORF2 | 5 306~6 940 | 5 355~6 995 | 5 079~6 731 |
| ORF3 | 6 940~7 566 | 6 995~7 630 | 6 731~7 510 |
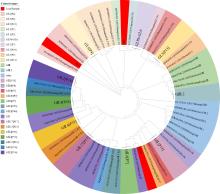
Figure 1
Phylogenetic tree based on the whole-genome sequences of NoV strains

Table 2
Nucleotide homology between the three NoV strains and their reference sequences in the ORF1, ORF2 and ORF3 regions(%)
| 毒株 | LC122690.1 | MW243610.2 | PV640830.1 | NR18 | NR246 | NR61 | ||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| ORF1 | ORF2 | ORF3 | ORF1 | ORF2 | ORF3 | ORF1 | ORF2 | ORF3 | ORF1 | ORF2 | ORF3 | ORF1 | ORF2 | ORF3 | ORF1 | ORF2 | ORF3 | |||||||
| LC122690.1 | 100.0 | 100.0 | 100.0 | 37.9 | 36.9 | 32.6 | 70.5 | 67.2 | 62.1 | 70.5 | 67.1 | 62.2 | 38.3 | 37.0 | 32.5 | 98.7 | 98.2 | 98.7 | ||||||
| MW243610 | 37.9 | 36.9 | 32.6 | 100.0 | 100.0 | 100.0 | 38.1 | 38.0 | 30.3 | 38.2 | 37.9 | 30.0 | 98.7 | 98.9 | 95.3 | 38.0 | 36.7 | 32.7 | ||||||
| PV640830.1 | 70.5 | 67.2 | 62.1 | 38.1 | 38.0 | 30.3 | 100.0 | 100.0 | 100.0 | 99.7 | 99.3 | 99.6 | 38.2 | 38.0 | 31.6 | 70.3 | 67.3 | 62.4 | ||||||
| NR18 | 70.5 | 67.1 | 62.2 | 38.2 | 37.9 | 30.0 | 99.7 | 99.3 | 99.6 | 100.0 | 100.0 | 100.0 | 38.3 | 37.8 | 31.4 | 70.3 | 67.3 | 62.5 | ||||||
| NR246 | 38.3 | 37.0 | 32.5 | 98.7 | 98.9 | 95.3 | 38.2 | 38.0 | 31.6 | 38.3 | 37.8 | 31.4 | 100.0 | 100.0 | 100.0 | 38.3 | 36.8 | 32.6 | ||||||
| NR61 | 98.7 | 98.2 | 98.7 | 38.0 | 36.7 | 32.7 | 70.3 | 67.3 | 62.4 | 70.3 | 67.3 | 62.5 | 38.3 | 36.8 | 32.6 | 100.0 | 100.0 | 100.0 | ||||||
Table 3
Amino acid homology of VP1 and VP2 regions between the three NoV strains and their reference sequences(%)
| 毒株 | LC122690.1 | MW243610.2 | PV640830.1 | NR18 | NR246 | NR61 | ||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| VP1 | VP2 | VP1 | VP2 | VP1 | VP2 | VP1 | VP2 | VP1 | VP2 | VP1 | VP2 | |||||||
| LC122690.1 | 100.0 | 100.0 | 12.9 | 11.8 | 51.5 | 51.5 | 51.2 | 52.0 | 13.1 | 11.6 | 96.3 | 97.5 | ||||||
| MW243610 | 12.9 | 11.8 | 100.0 | 100.0 | 12.4 | 10.4 | 12.2 | 10.0 | 97.9 | 93.5 | 12.4 | 11.5 | ||||||
| PV640830.1 | 51.5 | 51.5 | 12.4 | 10.4 | 100.0 | 100.0 | 98.8 | 99.5 | 12.6 | 11.4 | 52.2 | 52.0 | ||||||
| NR18 | 51.2 | 52.0 | 12.2 | 10.0 | 98.8 | 99.5 | 100.0 | 100.0 | 12.4 | 11.0 | 51.9 | 52.5 | ||||||
| NR246 | 13.1 | 11.6 | 97.9 | 93.5 | 12.6 | 11.4 | 12.4 | 11.0 | 100.0 | 100.0 | 12.6 | 11.2 | ||||||
| NR61 | 96.3 | 97.5 | 12.4 | 11.5 | 52.2 | 52.0 | 51.9 | 52.5 | 12.6 | 11.2 | 100.0 | 100.0 | ||||||
Table 4
Amino acid mutations in the VP1, VP2 regions between the NR18 strain and its reference sequence
| 突变位点 | 140 | 244 | 250 | 291 | 362 | 403 | 425 | 506 | 111* |
|---|---|---|---|---|---|---|---|---|---|
| NR18 | C | P | V | P | S | A | S | ||
| PV640830.1 | S | Q | S | I | S | P | Q | T | F |
Table 5
Amino acid mutations in the VP1, VP2 regions between the NR61 strain and its reference sequence
| 突变位点 | 52 | 64 | 79 | 86 | 125 | 136 | 141 | 157 | 194 | 203 | 205 | 210 | 226 | 229 | 249 | 254 | 291 | 294 | 295 | 331 | 332 | 364 | 398 | 406 | 411 | 496 | 517 | 81* | 100* | 125* | 129* | 152* | 160* | 168* | 175* |
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| NR61 | F | I | T | I | P | S | Q | V | S | A | T | F | P | L | L | V | I | L | I | I | S | G | T | C | L | I | P | D | R | V | T | S | C | P | |
| LC122690.1 | S | T | I | T | L | F | L | A | L | V | I | S | L | P | P | W | A | T | S | T | T | L | D | I | Y | P | T | L | G | C | G | I | F | Y | L |
Table 6
Amino acid mutations in the VP1, VP2 regions between the NR246 strain and its reference sequence
| 突变位点 | 16 | 31 | 105 | 109 | 144 | 152 | 176 | 294 | 312 | 317 | 352 | 366 | 409 | 419 | 451 | 42* | 92* | 152* | 178* | 216* | 230* | 246* | 259* |
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| NR246 | N | V | W | A | S | G | A | R | Q | G | I | Q | L | P | K | A | K | A | T | ||||
| MW243610.2 | S | A | C | V | L | V | V | Q | R | D | T | R | W | S | L | K | R | V | Q | V | A | E |
| [1] |
Bartsch SM, Lopman BA, Ozawa S, et al. Global economic burden of norovirus gastroenteritis[J]. PLoS One, 2016, 11(4):e0151219.
doi: 10.1371/journal.pone.0151219 URL |
| [2] |
Zhou HL, Zhen SS, Wang JX, et al. Burden of acute gastroenteritis caused by norovirus in China: a systematic review[J]. J Infect, 2017, 75(3):216-224.
doi: 10.1016/j.jinf.2017.06.004 URL |
| [3] |
Teunis PFM, Moe CL, Liu PB, et al. Norwalk virus: How infectious is it?[J]. J Med Virol, 2008, 80(8):1468-1476.
doi: 10.1002/jmv.21237 pmid: 18551613 |
| [4] |
Belliot G, Sosnovtsev SV, Mitra T, et al. In vitro proteolytic processing of the MD145 norovirus ORF1 nonstructural polyprotein yields stable precursors and products similar to those detected in Calicivirus-infected cells[J]. J Virol, 2003, 77(20):10957-10974.
pmid: 14512545 |
| [5] |
Arias A, Emmott E, Vashist S, et al. Progress towards the prevention and treatment of norovirus infections[J]. Future Microbiol, 2013, 8(11):1475-1487.
doi: 10.2217/fmb.13.109 pmid: 24199805 |
| [6] |
Green KY, Belliot G, Taylor JL, et al. A predominant role for Norwalk-like viruses as agents of epidemic gastroenteritis in Maryland nursing homes for the elderly[J]. J Infect Dis, 2002, 185(2):133-146.
pmid: 11807686 |
| [7] |
Conley MJ, McElwee M, Azmi L, et al. Calicivirus VP2 forms a portal-like assembly following receptor engagement[J]. Nature, 2019, 565(7739):377-381.
doi: 10.1038/s41586-018-0852-1 |
| [8] |
Kroneman A, Vega E, Vennema H, et al. Proposal for a unified norovirus nomenclature and genotyping[J]. Arch Virol, 2013, 158(10):2059-2068.
doi: 10.1007/s00705-013-1708-5 pmid: 23615870 |
| [9] |
Chhabra P, de Graaf M, Parra GI, et al. Updated classification of norovirus genogroups and genotypes[J]. J Gen Virol, 2019, 100(10):1393-1406.
doi: 10.1099/jgv.0.001318 pmid: 31483239 |
| [10] |
Lopman B, Gastañaduy P, Park GW, et al. Environmental transmission of norovirus gastroenteritis[J]. Curr Opin Virol, 2012, 2(1):96-102.
doi: 10.1016/j.coviro.2011.11.005 pmid: 22440972 |
| [11] |
Kendra JA, Tohma K, Parra GI. Global and regional circulation trends of norovirus genotypes and recombinants, 1995-2019: a comprehensive review of sequences from public databases[J]. Rev Med Virol, 2022, 32(5):e2354.
doi: 10.1002/rmv.v32.5 URL |
| [12] |
Eden JS, Tanaka MM, Boni MF, et al. Recombination within the pandemic norovirus GⅡ.4 lineage[J]. J Virol, 2013, 87(11):6270-6282.
doi: 10.1128/JVI.03464-12 URL |
| [13] |
Ahmed SM, Hall AJ, Robinson AE, et al. Global prevalence of norovirus in cases of gastroenteritis: a systematic review and meta-analysis[J]. Lancet Infect Dis, 2014, 14(8):725-730.
doi: S1473-3099(14)70767-4 pmid: 24981041 |
| [14] |
Lindsay L, Wolter J, de Coster I, et al. A decade of norovirus disease risk among older adults in upper-middle and high income countries: a systematic review[J]. BMC Infect Dis, 2015, 15:425.
doi: 10.1186/s12879-015-1168-5 pmid: 26467099 |
| [15] |
Hong XJ, Xue L, Cao YW, et al. The variation of antigenic and histo-blood group binding sites synergistically drive the evolution among chronologically emerging GⅡ.4 noroviruses[J]. Heliyon, 2024, 10(5):e26567.
doi: 10.1016/j.heliyon.2024.e26567 URL |
| [16] |
Karst SM, Baric RS. What is the reservoir of emergent human norovirus strains?[J]. J Virol, 2015, 89(11):5756-5759.
doi: 10.1128/JVI.03063-14 pmid: 25787285 |
| [17] | 史永林, 傅荔艳, 金晶, 等. 2020—2021年安徽省急性胃肠炎疫情中GⅠ组诺如病毒基因型多样性分析[J]. 国际病毒学杂志, 2022, 29(2):103-108. |
| [18] |
Hu XY, Sun LL, Han TL, et al. The genetic diversity of genogroup Ⅰ noroviruses causing acute gastroenteritis outbreaks in Beijing between 2014 and 2023[J]. Heliyon, 2024, 10(20):e39202.
doi: 10.1016/j.heliyon.2024.e39202 URL |
| [19] |
Arana A, Cilla G, Montes M, et al. Genotypes, recombinant forms, and variants of norovirus GⅡ.4 in Gipuzkoa (Basque Country, Spain), 2009-2012[J]. PLoS One, 2014, 9(6):e98875.
doi: 10.1371/journal.pone.0098875 URL |
| [20] |
Lim KL, Hewitt J, Sitabkhan A, et al. A multi-site study of norovirus molecular epidemiology in Australia and New Zealand, 2013-2014[J]. PLoS One, 2016, 11(4):e0145254.
doi: 10.1371/journal.pone.0145254 URL |
| [21] | Yu YX, Yan SL, Li BL, et al. Genetic diversity and distribution of human norovirus in China (1999-2011)[J]. BioMed Res Int, 2014, 2014:196169. |
| [22] | Dong XG, Qin M, Wang ZE, et al. Should we pay attention to recombinant norovirus strain GⅡ.P7/GⅡ.6?[J]. J Infect Public Heal, 2019, 12(3):403-409. |
| [23] | 王越, 郎兴莹, 于蕾, 等. 2020—2023年大连地区诺如病毒胃肠炎疫情分子流行特征分析[J]. 病毒学报, 2024, 40(4):857-861. |
| [24] | 陶璐秋, 王璇, 余艳, 等. 2019—2022年南京市急性胃肠炎暴发疫情中诺如病毒分子流行病学分析[J]. 病毒学报, 2023, 39(5):1272-1284. |
| [25] | 纪蕾, 刘光涛, 沈月华, 等. 2013—2019年浙江省湖州市急性胃肠炎病例GⅡ.P7-GⅡ.6型诺如病毒基因特征分析[J]. 疾病监测, 2021, 36(1):53-58. |
| [26] |
Allen DJ, Gray JJ, Gallimore CI, et al. Analysis of amino acid variation in the P2 domain of the GⅡ-4 norovirus VP1 protein reveals putative variant-specific epitopes[J]. PLoS One, 2008, 3(1):e1485.
doi: 10.1371/journal.pone.0001485 URL |
| [27] |
Tan M, Jiang X. The P domain of norovirus capsid protein forms a subviral particle that binds to histo-blood group antigen receptors[J]. J Virol, 2005, 79(22):14017-14030.
doi: 10.1128/JVI.79.22.14017-14030.2005 pmid: 16254337 |
| [28] |
Izquierdo-Lara RW, Villabruna N, Hesselink DA, et al. Patterns of the within-host evolution of human norovirus in immunocompromised individuals and implications for treatment[J]. eBioMedicine, 2024, 109:105391.
doi: 10.1016/j.ebiom.2024.105391 URL |
| [29] |
Lindesmith LC, Beltramello M, Donaldson EF, et al. Immunogenetic mechanisms driving norovirus GⅡ.4 antigenic variation[J]. PLoS Pathog, 2012, 8(5):e1002705.
doi: 10.1371/journal.ppat.1002705 URL |
| [30] |
Bull RA, Hansman GS, Clancy LE, et al. Norovirus recombination in ORF1/ORF2 overlap[J]. Emerg Infect Dis, 2005, 11(7):1079-1085.
pmid: 16022784 |
| [1] | DU Yanli, LIU Yaqiong, ZHOU Xingyu, YUAN Heng. Epidemiology and pathogen spectrum of other infectious diarrhea in Sichuan Province from 2019 to 2023 [J]. Journal of Tropical Diseases and Parasitology, 2026, 24(1): 7-12. |
| [2] | ZHENG Erda, JIA Yuchen, LIU Sixian, SHEN Xiulian, HUANG Tian, HE Jibo, LIU Huihui. Epidemiological characteristics and prediction of epidemic trend of norovirus infections in Yunnan Province [J]. Journal of Tropical Diseases and Parasitology, 2025, 23(4): 222-228. |
| [3] | YAN Wenzhe, LI Jinyu, LUN Xinchang, LIU Pengbo, SONG Xiuping, GUO Yuhong, LIU Xiaobo, YUE Yujuan, LU Liang, ZHAO Ning. Study on the virus diversity of the mosquitoes carrying Japanese encephalitis virus [J]. Journal of Tropical Diseases and Parasitology, 2024, 22(6): 328-333. |
| [4] | GE Yinglu, YANG Lingkang, LIU Yinuo, MA Wanwan, WANG Peng, SUN Yong, SHI Yonglin. Molecular characterization of coxsackievirus A16 associated hand, foot and mouth disease in Anhui Province from 2021 to 2023 [J]. Journal of Tropical Diseases and Parasitology, 2024, 22(4): 233-238. |
| [5] | LI Jin-song. Policies, current situation and challenge of prevention and control of viral diarrhea in China [J]. Journal of Tropical Diseases and Parasitology, 2023, 21(2): 61-64. |
| [6] | WANG Xue-fei, TIAN Rong, CHEN Zhi-hui, YANG Ming-dong, GUO Xiao-fang. Analysis on the epidemiological and etiological characteristics of imported dengue fever cases in Kunming in 2019 [J]. Journal of Tropical Diseases and Parasitology, 2022, 20(4): 215-220. |
| [7] | CUI Dong, XIAO Na, ZHONG Guan-nan, YANG Xiao-hua, GU Jia-ying, HONG Wen-teng. Investigation on the enterovirus and infectious diarrhea virus in population in Yantian district of Shenzhen [J]. Journal of Tropical Diseases and Parasitology, 2021, 19(6): 322-324,353. |
| [8] | RAO Tian-yu△, SHEN Xi△, ZHU Zhi-wei, XU Wei, ZHAO Jin-hong∗. Morphological and molecular characteristics of Ascaridia galli isolates from Huangshan area [J]. Journal of Tropical Diseases and Parasitology, 2020, 18(2): 87-92. |
| Viewed | ||||||
|
Full text |
|
|||||
|
Abstract |
|
|||||
